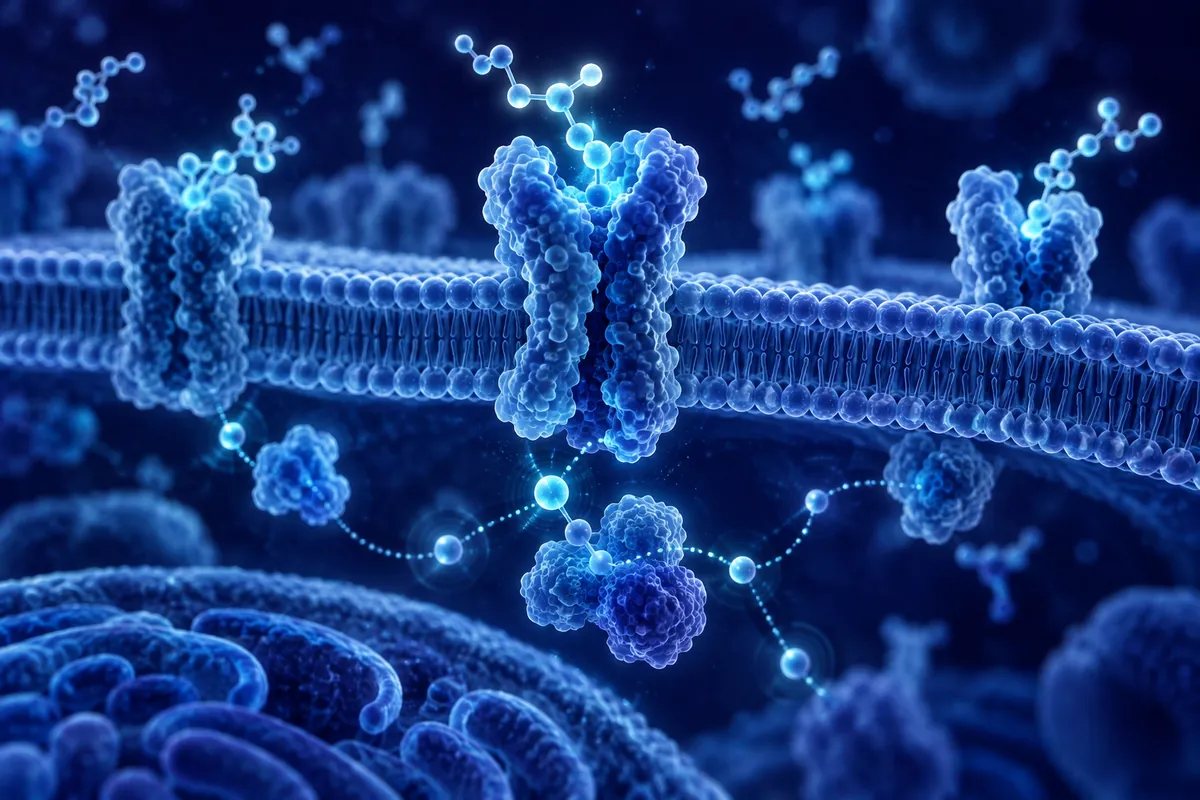
Peptide pharmacology has spent the last decade chasing polypharmacology : one molecule, several receptor targets, additive or synergistic signaling. Dual GLP-1/GIP agonists proved the concept could work cleanly in a single chain. Triple agonists added glucagon receptor activity on top of that backbone. Now a newer design class pushes further still: quintuple agonists that fuse incretin receptor engagement with a small-molecule PPAR ligand, delivering nuclear receptor pharmacology to cells that express GLP-1, GIP, or glucagon receptors on their surface. It’s an elegant piece of chemistry, and it raises genuinely interesting questions about targeted delivery that go beyond simple receptor counting.
From Receptor Agonism to Conjugate Chemistry
A classical incretin peptide binds a class B GPCR, triggers Gs-coupled cAMP signaling, and dissociates. A quintuple agonist construct is built differently. Researchers start with a multi-receptor peptide backbone : often a GLP-1/GIP/glucagon tri-agonist scaffold stabilized with an Aib substitution at position 2 to resist DPP-4 cleavage : and conjugate it, via a stable linker, to a nuclear receptor ligand such as a PPAR-alpha/gamma dual agonist. The linker chemistry matters enormously here. Amide or triazole linkages are common choices because they resist plasma esterases while still permitting intracellular release once the conjugate is internalized through receptor-mediated endocytosis.
Why go to this trouble instead of just co-administering two separate compounds in a rodent protocol? Because PPAR ligands, left unconjugated, distribute broadly across tissues expressing the receptor isoform : adipocytes, hepatocytes, and beyond : with no cell-type selectivity. Fusing the ligand to a peptide that only activates its own receptor on specific cell populations essentially converts a systemically acting nuclear receptor agonist into what the discovery literature calls a “cell-targeted” or “guided-missile” pharmacophore.
Nuclear Receptor Pharmacology, Delivered Selectively
PPAR-alpha and PPAR-gamma are ligand-activated transcription factors that heterodimerize with RXR and bind PPAR response elements in promoter regions, altering expression of genes involved in lipid handling and mitochondrial biogenesis. Unlike GPCRs, which signal in seconds, nuclear receptor activation unfolds over hours as transcriptional programs are induced. That mismatch in kinetics is part of the design challenge: a peptide conjugate must remain intact long enough for internalization and intracellular ligand release, yet the incretin-receptor arm still needs to preserve normal Gs/cAMP signaling kinetics at the cell surface. Structure-activity work in HEK293 cells stably expressing human GLP-1R, GIPR, and glucagon receptor constructs has been used to confirm that conjugation doesn’t blunt the peptide’s own binding affinity : measured by radioligand displacement or BRET-based cAMP assays : relative to the unconjugated parent peptide.
In rodent studies, this triple-plus-PPAR design has been examined primarily in diet-induced obese mouse models, where researchers track markers of hepatic lipid accumulation, adipose tissue gene expression, and energy expenditure via indirect calorimetry. No claims about outcomes in humans are appropriate here, and none are made in the primary literature describing these constructs : the translational gap between rodent adipose biology and human metabolic physiology remains wide, and species differences in PPAR isoform expression ratios are a recurring caveat investigators flag when interpreting rodent data.
The “Exercise Mimetic” Framing
You’ll sometimes see these conjugates described informally as exercise mimetics. The phrase refers to a mechanistic overlap, not a functional promise: chronic aerobic exercise activates PGC-1alpha and downstream PPAR-delta/gamma transcriptional programs in skeletal muscle and adipose tissue, and some quintuple-agonist constructs appear to engage overlapping transcriptional nodes in rodent tissue. That’s a mechanistic parallel worth studying, not evidence that a molecule replicates everything exercise does physiologically. Researchers working with these conjugates in cell culture typically use C2C12 myotube models or primary adipocyte cultures to probe which transcriptional programs are actually induced, using RT-qPCR panels for genes like PDK4, UCP1, and CPT1A rather than relying on the mimetic label as a stand-in for mechanistic data.
Assay Considerations for Multi-Target Constructs
Characterizing a five-way pharmacophore is not trivial. Each receptor arm needs its own selectivity and potency data, and cross-reactivity has to be ruled out systematically. A typical panel might include: cAMP accumulation assays across GLP-1R, GIPR, and glucagon receptor-transfected cell lines to confirm agonist activity at each incretin target; a PPAR transactivation reporter assay, often a luciferase construct driven by a PPRE, to quantify nuclear receptor engagement; and LC-MS/MS-based stability testing in plasma or liver microsomes to characterize linker cleavage kinetics. Binding kinetics are frequently profiled by surface plasmon resonance to distinguish genuine dual-mode engagement from artifacts of assay interference. None of this is optional : a conjugate that looks potent in a single cAMP assay but degrades within minutes in microsomal preparations isn’t a viable lead, however clean its initial binding data look.
Where the Research Questions Sit Now
Quintuple agonism sits at an unusual intersection of GPCR pharmacology and nuclear receptor biology, and the open questions are as much about chemistry as about biology. How much PPAR ligand payload can a peptide carry before pharmacokinetics degrade? Does intracellular ligand release actually achieve meaningfully higher local PPAR occupancy than systemic dosing would in equivalent rodent tissue, or does the conjugate simply redistribute exposure without changing total transcriptional output? And how do investigators disentangle GPCR-driven acute signaling from the slower nuclear receptor transcriptional response when both pathways converge in the same cell? These are the questions occupying medicinal chemistry and pharmacology groups working on multi-target peptide-small-molecule conjugates, and in vitro and rodent in vivo models remain the appropriate tools for answering them at this stage of investigation.
Disclaimer: This content is intended for research purposes only and is not meant to constitute medical advice.
This educational material discusses laboratory research concepts only. Loti Labs products are intended strictly for research and development use and are not for human consumption.